Practical Protein Chemistry - A. Darbre 1989
Determination of the composition of protein oligomers. Isolation of monomers and polypeptide chains
Cross-linking of oligomer and monomer
Disulfide bonds
Data from N-terminal Amino acid analysis provide Evidence for the cross-linking of individual subunits. For example, a yield of an N-terminal amino acid greater than 1 mol/mol of protein can indicate the presence of two identical polypeptide chains in the oligomer. Similarly, an N-terminal amino acid yield (excluding Serine) of around 50% may suggest that the N-terminal amino acid of one of the two chains is blocked. Because the yield of an N-terminal amino acid depends on numerous factors, premature Conclusions should be avoided when interpreting these results.
Various types of cross-links have been identified in Proteins: disulfide, aldol, and aldimine bonds. Among these, S—S bonds are the most labile, which is why disulfide Cleavage is most frequently employed in practice. The resulting polypeptide chains are separated using standard Methods established for monomer Separation (Section 1.3.2).
Disulfide Bonds are cleaved via oxidation, reduction, or Nucleophilic substitution reactions (HSO-3, CN-, BH-4, H-) [40, 106] (see also Chapter 2). Oxidation of S—S bonds yields two cysteic acid residues [160]. Additional sulfo groups impart significant hydrophilicity to the protein molecule, enhancing its Water solubility, particularly at low pH. However, The oxidation of disulfide bonds is accompanied by the modification of Other Amino Acids and partial Hydrolysis of labile peptide bonds. Therefore, to introduce sulfo groups, reductive Cleavage of disulfide Bonds followed by mild oxidation of SH groups using sultones (internal sulfonic acid esters) is preferred (see Sections 1.5.1.2 and 2.2.2.3).
Disulfide bonds are typically cleaved by reduction, and the reaction progress is monitored using Ellman's reagent [69, 102]. Following reduction, the generated SH groups are alkylated to prevent re-oxidation and cystine formation. The alkylating agent is chosen based on the specific experimental goals—for instance, to increase Protein solubility in aqueous buffers, introduce additional cleavage sites for Trypsin, or introduce a labile protecting group for subsequent regeneration of SH groups. In some cases, SH protection may be unnecessary, as proteins with reduced S—S bonds can remain stable in cold acetic acid at pH 3 [7].
1.5.1.1. Cleavage of Disulfides.
Oxidation of Cysteine or disulfide bonds in proteins is recommended to be carried out using performic acid [78] (see also Section 2.2.3.1).
After oxidation, the Amino Acid Composition of the test protein is determined, with a primary focus on cysteic acid content. The yield of cysteic acid should be ≥85%, and that of Methionine S,S-dioxide (if methionine is present) at least 95%; Tryptophan is completely destroyed during oxidation. In the presence of chlorides, Tyrosine modification may occur; therefore, it is advisable to remove chloride ions from the sample prior to analysis. A Column of Dowex 2 (100 mesh) in the acetate form is washed with 3 M sodium acetate until chloride ions are absent in the eluate (monitored by the silver nitrate test). The protein solution (0.1 g of protein per 1 ml of ion-exchanger bed volume) is then applied to the column and washed with a small volume of buffer, taking into account potential protein loss on the column.
Reduction of S—S bonds is carried out primarily using 2-mercaptoethanol. However, if used in insufficient excess, 2-mercaptoethanol can form mixed disulfides and has an unpleasant odor. Dithiothreitol and dithioerythritol are highly effective alternatives that ensure complete reduction even in small excess. In several studies, disulfide bonds have been reduced using tributylphosphine [150–152] (these methods are discussed in more detail in Section 2.2.1).
Disulfide bonds linking two or more polypeptide chains are generally accessible to Solvents and Reagents even in the absence of Denaturing Agents, whereas intrachain S—S bonds are typically masked and inaccessible. This allows for the Selective reduction of exposed S—S bonds. Complete reduction of disulfides is recommended to be performed in a mildly acidic medium in the presence of urea or guanidine hydrochloride. None of the denaturing agents are entirely free of drawbacks; for instance, trace urea in a sample generates cyanate ions during the Condensation step of Edman Degradation, which can interfere with sequencing. Experience with guanidine-HCl is less problematic, and when a well-purified preparation is used, sequencing issues are generally not observed.
In studies on yG-immunoglobulin, a method was developed to reduce interchain S—S bonds while preserving intrachain ones [53].
The resulting fragments are analyzed for SH-group content, or the mixture is fractionated in the presence of reducing agents or under acidic conditions. For convenience, thiol-containing fragments are converted into more stable or more soluble derivatives.
Carboxymethylation is the most thoroughly studied yet imperfect method for SH-group protection [68]. It should be noted that if The conversion of thiazolinones into phenylthiohydantoins during Edman sequencing is performed under insufficiently mild conditions, significant losses of S-carboxymethylcysteine may occur [84]. Alkylation with 14C-iodoacetamide is a useful approach, though Other reagents are also successfully employed.
If subsequent protein refolding studies are planned, derivatives with labile protecting groups can be prepared. Some recommended reagents require careful handling; for example, iodoacetic acid is an allergen, and work with this reagent is typically performed using rubber gloves in a well-ventilated area (see also Chapter 2).
1.5.1.2. Reversible Modification. The action of sulfite on cystine yields cysteine and cysteine sulfonate (sulfocysteine) (see also Section 2.2.4).
Class="center">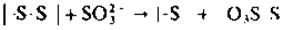
Disulfide bonds can be reconstructed by reducing the modified protein and refolding it in the presence of atmospheric oxygen. Oxidative sulfitolysis of disulfides in the presence of copper salts leads to The formation of S-sulfonates [37].
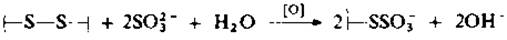
Similar results were obtained by reacting a protein with sodium sulfite in 8 M urea in the presence of atmospheric oxygen and traces of cysteine [31]. Reduction of cystine with dithiothreitol followed by Treatment with sodium tetrathionate yields S-sulfocysteine quantitatively [85]. S-Sulfocysteine is stable at neutral pH and under Edman degradation conditions, but reverts to cysteine upon reduction. Protein Hydrolysis with 6 M HCl converts S-sulfocysteine into cystine [37]. Monitoring reaction completion presents a significant challenge; when using [35S]sulfite, incorporation of the label is used for tracking [31].
The reaction of w-toluenesultone with a reduced protein at pH 8.3 yields S-2-sulfobenzylcysteine [153].
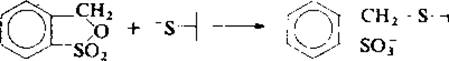
The reverse reaction is carried out by reduction with sodium in liquid ammonia; sodium amide is added to the reaction mixture to prevent the cleavage of Thr-Pro bonds. Proteins modified via sulfitolysis and sulfobenzylation are readily soluble in water.
Oxidative sulfitolysis [31]. This method has been successfully applied in studies of aldolase, Lactate dehydrogenase, and pepsinogen. Reaction completion is achieved only within the pH range of 7.0–8.5, indicating the involvement of the protonated amino group of cysteine. A protein sample (2 mg/ml) is incubated in the presence of atmospheric oxygen at 25 °C for 1 h in a solution of the following composition: 0.1 M Tris-HCl (pH 8.4) + 8 M urea (or 6 M guanidine-HCl) + 0.05 M sodium sulfite, 0.2 mM cysteine. Upon completion, the reaction mixture is dialyzed and lyophilized.
Following sulfitolysis, the active enzyme (aldolase) is obtained using the following Procedure. A protein sample (0.5–1 mg/ml) is dissolved in 0.1 M Tris-HCl (pH 7.5) containing 4 M urea, 2-mercaptoethanol is added (to a final concentration of 0.7 mol/l), and the mixture is incubated at 25 °C for 2 h. The reaction mixture is then diluted 10-fold with a buffer solution of the following composition: 0.1 M Tris-HCl (pH 7.5) + 2.5 mg/ml bovine serum albumin + 25 mM EDTA + 10 mM 2-mercaptoethanol; it is incubated in the presence of atmospheric oxygen, and after 15 min, The activity of the refolded enzyme is measured.
S-Sulfobenzylation [153]. A protein sample (100 µmol) in 100 ml of 0.5 M sodium bicarbonate mixed with propanol (1:1) is incubated under a nitrogen atmosphere with tributylphosphine (330 µmol) and w-toluenesultone (204 mg, 1200 µmol). Excess w-toluenesultone is removed by filtration, the filtrate is concentrated to a residual volume of 40 ml, and dialyzed against water. On an amino acid analyzer, S-sulfobenzylcysteine elutes immediately after cysteic acid. When reacting with Insulin, the yield of cysteine derivatives is quantitative.
Removal of the S-sulfobenzyl group [90]. 300 mg of protein is thoroughly dried and dissolved in anhydrous liquid ammonia (300 ml) in a round-bottomed flask cooled with a dry ice-acetone bath. To protect the labile peptide bonds of Proline residues, ~120 mg of sodium amide is added to the reaction mixture. Subsequent steps are carried out at the boiling point of liquid ammonia. A freshly prepared solution of metallic sodium in liquid ammonia is added dropwise until the reaction mixture acquires a faint blue color. After 30 s, a few drops (2–3) of glacial acetic acid are added until the color disappears, and the ammonia is evaporated to a residual volume of 10 ml. The residue is concentrated in vacuo on a rotary evaporator and dried to completion using a water aspirator.
1.5.1.3. Oxidation. The reconstruction of most disulfide bonds in certain proteins is achieved by atmospheric oxidation, i.e., by aerating a solution of the reduced protein [70]. Apparently, the native protein molecule possesses a thermodynamically most favorable conformation, and therefore, the primary factor driving the "correct" approximation of thiol groups is the cooperative interaction of the functional groups within the polypeptide chain. If the distribution of disulfide bonds in a protein is mainly determined by the conformation of its precursor (e.g., proinsulin in the case of insulin), such proteins refold with difficulty. Below is a standard procedure for protein oxidation aimed at renaturation; however, optimal reaction conditions must be determined empirically based on the individual Properties of the protein under study, its Primary Structure, and the three-dimensional folding of the polypeptide chain. For instance, The addition of 2-mercaptoethanol and an increase in Temperature to 38 °C proved beneficial for the renaturation of Lysozyme, but ineffective for Ribonuclease [56]. For other proteins, the optimal concentration of 2-mercaptoethanol is selected empirically. Both factors—low protein concentration and the presence of 2-mercaptoethanol—prevent subunit aggregation and the formation of intermolecular disulfide bonds. It is hypothesized that 2-mercaptoethanol catalyzes the disulfide exchange required to correct mispaired cysteine residues.
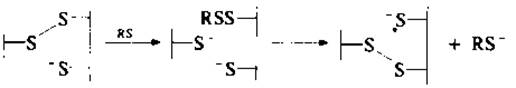
Metal Ions inhibit activation due to binding with thiols. Activation involves the slow dialysis of denaturing agents, such as urea. Carrying out the reaction in open vessels is entirely sufficient for oxygen access; vigorous aeration is not recommended as it may cause Protein Denaturation.
Procedure 1 [56]. The renaturation of reduced lysozyme (0.025 mg/mL) is carried out in 0.1 M Tris-buffer (pH 8.5) at 38 °C for 2 h in the presence of 2-mercaptoethanol (at a molar ratio of 2-mercaptoethanol : cysteine = 180 : 1). The yield of the activated product is 70%.
Procedure 2 [52]. Reduced insulin chains (5 mg protein/mL) are precipitated at pH 3.8 and thoroughly washed with deaerated buffer in the cold. Atmospheric oxidation is performed in Glycine buffer (pH 10.6) at 35 °C. The renatured protein exhibits 50% activity compared to the crystalline preparation. The activity of the resulting hormone depends on the relative content of polypeptide chains in the reaction mixture; the maximum yield is achieved at an A:B ratio of 1.5:1.
Last update: 06/08/2026
Editorial and Educational Adaptation: This material has been compiled based on the primary/original source text. The project team performed an editorial review, corrected technical inaccuracies, structured sections, and adapted the content for an educational format.
What was processed:
- elimination of formatting defects (OCR errors, structural breaks, corrupted characters);
- editorial organization of content;
- standardization of terminology in accordance with academic sources;
- verification of factual statements against the original source text.
All mentions of the author, publication year, and origin of the primary text have been preserved in accordance with the source.