Orthopedics - Oleksa A.P. 2006
Pathophysiology of Joints
Developmental Defects and Congenital Skeletal Anomalies
Hypochondroplasia
Hypochondroplasia is an autosomal dominant inherited disorder with clinical manifestations similar to mild Achondroplasia, yet the skeletal Dysplasia presents with much milder features: affected individuals are taller than those with achondroplasia; the HEAD, facial features, and hands are normal; and there are no neurological manifestations. Lumbar lordosis and limb bowing are less pronounced (Specht E.E., Daentl D.L., 1975). These same authors note that hypochondroplasia is more frequently observed in females due to a higher rate of spontaneous Mutations and marked mental retardation.
Radiological examination reveals premature closure of the metaphyso-epiphyseal fusion of the long tubular bones with widened epiphyses, but with a normal growth plate. The feet exhibit a varus position resulting from the elongation of the fibulae. The interpedicular spaces of the spine are narrowed, though not to the same extent as in achondroplasia, and thus neurological symptoms are absent (Beals R.K., 1969) (Fig. 59). These radiological signs are less pronounced in early childhood and manifest somewhat later. Shortened limb segments can be lengthened via osteotomy and distraction osteogenesis using the Ilizarov method.
Class="center">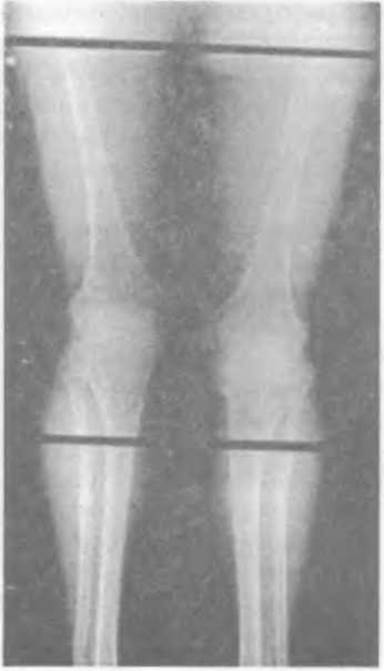
Fig. 59. Hypochondroplasia of the femurs in a 16-year-old girl (Deborah F. Bell K., 1993).
Last update: 10/08/2026
Editorial and Educational Adaptation: This material has been compiled based on the primary/original source text. The project team performed an editorial review, corrected technical inaccuracies, structured sections, and adapted the content for an educational format.
What was processed:
- elimination of formatting defects (OCR errors, structural breaks, corrupted characters);
- editorial organization of content;
- standardization of terminology in accordance with academic sources;
- verification of factual statements against the original source text.
All mentions of the author, publication year, and origin of the primary text have been preserved in accordance with the source.